预约演示
更新于:2026-09-12
Semaglutide (Zhongshan Wanhan)
司美格鲁肽(中山万汉)
更新于:2026-09-12
概要
基本信息
原研机构 |
非在研机构- |
权益机构- |
最高研发阶段临床1期 |
首次获批日期- |
最高研发阶段(中国)临床1期 |
特殊审评- |
登录后查看时间轴
关联
1
项与 司美格鲁肽(中山万汉) 相关的临床试验CTR20250353
随机、开放、单次给药、平行对照评价司美格鲁肽注射液与诺和泰在中国健康受试者中的药代动力学比对研究
100 项与 司美格鲁肽(中山万汉) 相关的临床结果
登录后查看更多信息
100 项与 司美格鲁肽(中山万汉) 相关的转化医学
登录后查看更多信息
100 项与 司美格鲁肽(中山万汉) 相关的专利(医药)
登录后查看更多信息
778
项与 司美格鲁肽(中山万汉) 相关的文献(医药)2026-12-31JOURNAL OF OBSTETRICS AND GYNAECOLOGY
Comparison of lifestyle, surgery, and semaglutide for weight management in endometrial cancer: a prospective observational study
Article
作者: Xiong, Yingchao ; Gong, Yuanyuan ; Li, Xiaodan ; Wei, Yuxuan ; Zhong, Zhaohui ; Gong, Jingjing ; Zeng, Hui
BACKGROUND:
To compare sleeve gastrectomy, semaglutide therapy, and the Diet, Exercise, Accompany and Refresh (DEAR) weight management programme for weight control, metabolic improvement, and oncological outcomes in patients with endometrial cancer receiving fertility-sparing treatment.
METHODS:
In this prospective observational study, 53 patients received sleeve gastrectomy (n = 10), semaglutide therapy (n = 23), or the DEAR programme (n = 20) after multidisciplinary assessment and shared decision-making. Anthropometric and metabolic measures and tumour response were assessed over two years.
RESULTS:
At 1 year, all groups lost weight; the sleeve gastrectomy group had the largest body mass index reduction compared with the semaglutide group (difference = 4.04 kg/m2, p < 0.05) and the DEAR group (difference = 2.12 kg/m2, p < 0.05). By 2 years, weight regain was less frequent in the DEAR group (14.3%) than in the semaglutide (55.6%) and sleeve gastrectomy (66.7%) groups (p = 0.037). The DEAR group showed more stable metabolic improvements and the highest complete tumour remission rate (85.0%), followed by the semaglutide group (73.9%) and the sleeve gastrectomy group (60.0%).
CONCLUSION:
Sleeve gastrectomy produced rapid short-term weight loss, whereas the DEAR programme showed better medium-term weight maintenance, metabolic stability, and tumour response. Integrated lifestyle support may be a useful medium-term strategy for patients seeking fertility preservation.
2026-12-31ANNALS OF MEDICINE
GLP-1 receptor agonists in stroke prevention: a narrative review on emerging therapeutic frontiers
Review
作者: Chikatimalla, Rahul ; Aggarwal, Kanishk ; Shah, Twinkle ; Perry, Griffin ; Shah, Ashka ; Banker, Himanshi ; Jain, Rohit
OBJECTIVES:
To evaluate the current evidence supporting the cerebrovascular protective effects of glucagon-like peptide-1 receptor agonists (GLP-1RAs) in individuals with type 2 diabetes mellitus (T2DM), and to outline their mechanisms of action in stroke prevention.
METHODS:
A narrative review was conducted by synthesising data from cardiovascular outcome trials, meta-analyses and mechanistic studies involving GLP-1RAs such as semaglutide, liraglutide and dulaglutide. The search included literature on ischaemic stroke incidence, molecular pathways and clinical outcomes associated with GLP-1RA therapy.
RESULTS:
GLP-1RAs exhibit multiple protective mechanisms, including anti-inflammatory, antioxidant, neuroprotective and endothelial-stabilising effects. Long-acting agents demonstrate superior efficacy in reducing nonfatal and ischaemic stroke risk, with relative risk reductions ranging from 15% to 39% across major trials. These benefits are observed independent of glycemic control and appear most prominent in patients with preserved renal function and shorter diabetes duration. In contrast, short-acting exendin-based GLP-1RAs show limited cerebrovascular benefit. Treatment response may vary based on factors such as stroke subtype, baseline vascular risk and comorbidities.
CONCLUSION:
GLP-1RAs offer significant promise as adjunctive pharmacotherapy for stroke prevention in individuals with T2DM. Their multifactorial benefits extend beyond glucose regulation and may influence clinical outcomes through systemic vascular and neuroprotective mechanisms. However, inconsistencies in trial outcomes and limited data in non-diabetic or high-risk populations underscore the need for targeted stroke-specific studies. Personalised treatment approaches and broader risk stratification may optimise their use in cerebrovascular disease management.
2026-11-01FREE RADICAL BIOLOGY AND MEDICINE
Semaglutide ameliorates traumatic sepsis-induced cardiac injury by activating the March5-ACSL4 axis to suppress ferroptosis
Article
作者: Zhou, Feihu ; Shen, Yan ; Liu, Lingxin ; Sun, Baisheng ; Liu, Hui ; Kang, Hongjun ; Zhang, He ; Chen, Lanlan ; Wang, Xiaoli
Post-traumatic septic myocardial dysfunction (PT-SIMD) is a life-threatening condition with high mortality and no specific treatment. Ferroptosis, an iron-dependent regulated cell death pathway driven by lipid peroxidation, has been implicated in septic cardiac injury, yet the endogenous mechanisms that counteract this process remain poorly understood. We aimed to identify novel cardioprotective regulators of ferroptosis in PT-SIMD using unbiased proteomics and to investigate whether the GLP-1 receptor agonist semaglutide (SEMA) confers protection through modulation of mitochondrial E3 ubiquitin ligases. In a rat model combining limb crush injury and cecal ligation and puncture (CLP), PT-SIMD was accompanied by marked myocardial ferroptosis, evidenced by elevated ACSL4, reduced GPX4/FSP1, and 4-HNE accumulation. Quantitative 4D-DIA proteomic profiling identified March5, a mitochondrial E3 ubiquitin ligase, as significantly downregulated. In primary neonatal rat ventricular myocytes, LPS exposure suppressed March5 expression in a dose- and time-dependent manner. Knockdown of March5 using siRNA exacerbated LPS-induced ferroptosis, increasing ACSL4 protein levels, labile Fe2+, and lipid peroxidation. Notably, SEMA treatment upregulated March5 expression, attenuated ACSL4 accumulation, and ameliorated ferroptotic injury. Crucially, siRNA-mediated silencing of March5 in cardiomyocytes completely abolished the anti-ferroptotic and cytoprotective effects of SEMA. Collectively, March5 functions as a critical endogenous suppressor of ferroptosis in PT-SIMD by negatively regulating ACSL4. Semaglutide exerts its cardioprotective action via March5-dependent inhibition of the ACSL4 ferroptosis axis, highlighting a promising therapeutic strategy for sepsis-associated myocardial dysfunction.
100 项与 司美格鲁肽(中山万汉) 相关的药物交易
登录后查看更多信息
外链
| KEGG | Wiki | ATC | Drug Bank |
|---|---|---|---|
| - | - | - |
研发状态
10 条进展最快的记录, 后查看更多信息
登录
| 适应症 | 最高研发状态 | 国家/地区 | 公司 | 日期 |
|---|---|---|---|---|
| 2型糖尿病 | 临床1期 | 中国 | 2025-02-06 | |
| 2型糖尿病 | 临床1期 | 中国 | 2025-02-06 | |
| 肥胖 | 临床申请批准 | 中国 | 2025-12-26 |
登录后查看更多信息
临床结果
临床结果
适应症
分期
评价
查看全部结果
登录后查看更多信息
转化医学
使用我们的转化医学数据加速您的研究。
登录
或

药物交易
使用我们的药物交易数据加速您的研究。
登录
或

核心专利
使用我们的核心专利数据促进您的研究。
登录
或

临床分析
紧跟全球注册中心的最新临床试验。
登录
或

批准
利用最新的监管批准信息加速您的研究。
登录
或

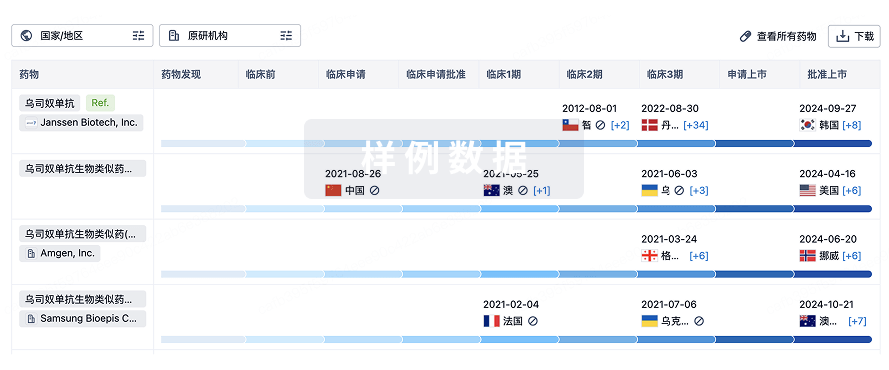
生物类似药
生物类似药在不同国家/地区的竞争态势。请注意临床1/2期并入临床2期,临床2/3期并入临床3期
登录
或
特殊审评
只需点击几下即可了解关键药物信息。
登录
或

提出任何生物医药研究问题
让每一步生物医药决策都精准制胜
基于十亿级生物数据与全球情报的精准 AI,赋能早期药物发现
立即开始免费试用!
智慧芽新药情报库是智慧芽专为生命科学人士构建的基于AI的创新药情报平台,助您全方位提升您的研发与决策效率。
立即接入生物医药数据包
智慧芽新药库数据也通过智慧芽开放平台,以MCP或API服务形式对外开放。立即使用我们的MCP服务定制您的LLM智能体。
生物序列数据库
生物药研发创新
免费使用
化学结构数据库
小分子化药研发创新
免费使用