预约演示
更新于:2026-10-19
Obinutuzumab
奥妥珠单抗
更新于:2026-10-19
概要
基本信息
药物类型 单克隆抗体 |
别名 Afutuzumab、Anti-CD20 monoclonal antibody R7159、Gazyvaro + [22] |
靶点 |
作用方式 抑制剂 |
作用机制 CD20抑制剂(B淋巴细胞抗原CD20抑制剂)、ADCC(抗体依赖的细胞毒作用)、CD20定向的溶细胞作用 |
在研适应症 |
非在研适应症 |
原研机构 |
最高研发阶段批准上市 |
首次获批日期 美国 (2013-11-01), |
最高研发阶段(中国)批准上市 |
特殊审评孤儿药 (美国)、孤儿药 (欧盟)、孤儿药 (澳大利亚)、优先审评 (美国)、优先审评 (中国)、突破性疗法 (美国) |
登录后查看时间轴
结构/序列
Sequence Code 68713L
来源: *****
Sequence Code 9699891H
来源: *****
关联
373
项与 奥妥珠单抗 相关的临床试验NCT07847957
A Phase 1b and 2 Study of Roginolisib (IOA-244) Combined With Obinutuzumab Followed by Venetoclax in Patients With Refractory/Relapsed Chronic Lymphocytic Leukemia (CLL) After Progression on Prior Bruton Tyrosine Kinase (BTK) Inhibitors
NCT07674810
A Phase 2 Trial Of The Combination Of Zanubrutinib, Sonrotoclax, And Obinutuzumab For Patients With Treatment-Naïve Chronic Lymphocytic Leukemia
NCT07634289
Radiotherapy in Combination With Glofitamab in Relapsed/Refractory Diffuse Large B Cell Lymphoma: The Phase II Radio-GLO Study
100 项与 奥妥珠单抗 相关的临床结果
登录后查看更多信息
100 项与 奥妥珠单抗 相关的转化医学
登录后查看更多信息
100 项与 奥妥珠单抗 相关的专利(医药)
登录后查看更多信息
1,186
项与 奥妥珠单抗 相关的文献(医药)2026-12-31mAbs
Impact of afucosylation strategy on antibody function: a comparative study of glycoengineered anti-CD20 antibodies Obinutuzumab and Obinutuzumab beta
Article
作者: Sun, Mingze ; Huang, Shuo ; Shuang, Qing ; Sun, Jicui ; Shen, Qunfang ; Jiang, Lingling ; Li, Feng ; Yang, Lijing ; Zhang, Lunfeng ; Li, Jiangmei ; Chen, Hong ; Lin, Guangzhong
Enhancing antibody-dependent cellular cytotoxicity (ADCC) via N-glycan afucosylation of Asn-297 is a validated strategy to improve the clinical efficacy of therapeutic antibodies. However, the impact of distinct glycoengineering approaches on the function of antibodies has not been systematically elucidated. Here, we experimentally compared two type II anti-CD20 antibodies, Obinutuzumab (Gazyva®) and Obinutuzumab beta (MIL62, Bejescin®), which share identical amino acid sequences but exhibit divergent glycosylation profiles. MIL62 was engineered with complete core afucosylation (fucose < 0.1%) lacking bisecting N-acetylglucosamine (GlcNAc) via knockout of the GDP-fucose transporter (GFT) in Chinese hamster ovary (CHO) cells used to produce the antibody. Conversely, Gazyva was produced in CHO cells that overexpress β-1,4-N-acetylglucosaminyltransferase III (GnT-III) and α-mannosidase II (α-ManII), resulting in ~50% fucose content and >80% bisecting GlcNAc occupancy. These distinct glycoengineering strategies led to disparate functional outcomes: afucosylated MIL62 showed improved FcγRIIIA binding and ADCC potency, while Gazyva with bisecting GlcNAc modifications exhibited higher glycoform heterogeneity and reduced thermal stability. Both antibodies displayed comparable FcRn binding, mannosylation, sialylation, and murine pharmacodynamics, mediating complete depletion of B cells in blood, lymph nodes, and spleen. Upon antigen rechallenge, MIL62 suppressed specific antibody titers, indicating memory B-cell eradication and profound potential to prevent autoimmune relapse. This study demonstrates that distinct glycoengineering strategies fundamentally reshape the antibody glycan profile beyond merely reducing fucose. These structural differences not only fine-tune ADCC potency, but also impact antibody stability. Particularly, complete afucosylation of MIL62 led to optimized ADCC potency and effectively suppressed pathogenic B cells repopulation in a delta-like ligand 3 (DLL3) re-challenge model, providing a critical framework for designing next-generation antibodies with superior therapeutic efficacy.
2026-12-01PEDIATRIC NEPHROLOGY
Which patients should switch from rituximab to obinutuzumab in refractory nephrotic syndrome?
Letter
作者: Saito, Aya ; Fujinaga, Shuichiro ; Ono, Takahiro
2026-12-01PEDIATRIC NEPHROLOGY
Evaluation of the comparative efficacy and safety of obinutuzumab and other follow-up strategies after rituximab failure in refractory nephrotic syndrome
Article
作者: Tu, Juan ; Wan, Ling ; Zha, Haoyu ; Chen, Chaoying
BACKGROUND:
Refractory nephrotic syndrome (RNS) in children remains difficult to manage, particularly in patients who fail to maintain sustained remission after rituximab (RTX). Optimal follow-up treatment strategies after RTX failure remain unclear. This study aimed to evaluate the efficacy of different follow-up treatment strategies after RTX failure.
METHODS:
This retrospective study included children with RNS who failed to maintain sustained remission after a single course of RTX. According to subsequent treatment, patients were divided into three groups: anti-CD20-free maintenance therapy (ACF), RTX retreatment (RTX-R), and obinutuzumab treatment (OBZ). Patients in the ACF group received corticosteroids and/or other immunosuppressants without additional anti-CD20 biologics, whereas those in the RTX-R and OBZ groups received RTX or OBZ therapy respectively.
RESULTS:
Twenty-eight children were included: 10 in the ACF group, 8 in the RTX-R group, and 10 in the OBZ group. At 6 months, the OBZ group demonstrated significantly higher relapse-free survival compared to both the RTX-R group (100% vs. 37.5%, P < 0.01) and the ACF group (100% vs. 30%, P < 0.01), with no difference between the RTX and ACF groups. At 52 weeks, relapse-free survival remained highest in the OBZ group (80.0%), compared with 25.0% in the RTX-R group and 30.0% in the ACF group. OBZ induced sustained peripheral B-cell depletion, with B cells remaining undetectable throughout the scheduled 6-month follow-up. No serious adverse effects were observed in three groups.
CONCLUSION:
Obinutuzumab may be an effective and safe follow-up option for children with RNS who fail to maintain sustained remission after RTX.
1,461
项与 奥妥珠单抗 相关的新闻(医药)2026-10-18
抗体药物偶联物临床结果
2026-10-17
100 项与 奥妥珠单抗 相关的药物交易
登录后查看更多信息
研发状态
批准上市
10 条最早获批的记录, 后查看更多信息
登录
| 适应症 | 国家/地区 | 公司 | 日期 |
|---|---|---|---|
| 肾病综合征 | 美国 | 2026-09-25 | |
| 狼疮性肾炎 | 美国 | 2025-10-17 | |
| CD20阳性B细胞慢性淋巴细胞白血病 | 日本 | 2022-12-23 | |
| CD20阳性滤泡性淋巴瘤 | 日本 | 2018-07-02 | |
| CD20阳性滤泡性淋巴瘤 | 日本 | 2018-07-02 | |
| 滤泡性淋巴瘤 | 澳大利亚 | 2014-05-15 | |
| 慢性淋巴细胞白血病 | 美国 | 2013-11-01 |
未上市
10 条进展最快的记录, 后查看更多信息
登录
| 适应症 | 最高研发状态 | 国家/地区 | 公司 | 日期 |
|---|---|---|---|---|
| 特发性膜性肾病 | 申请上市 | 中国 | 2026-06-04 | |
| 特发性膜性肾病 | 申请上市 | 中国 | 2026-06-04 | |
| 特发性膜性肾病 | 申请上市 | 中国 | 2026-06-04 | |
| 先天性肾病 | 申请上市 | 日本 | 2026-05-14 | |
| 系统性红斑狼疮 | 申请上市 | 美国 | 2026-04-21 | |
| 边缘区B细胞淋巴瘤 | 申请上市 | 中国 | 2019-09-28 | |
| 膜性肾小球肾炎 | 临床3期 | 荷兰 | 2025-10-01 | |
| 肿瘤溶解综合征 | 临床3期 | 美国 | 2024-08-05 | |
| 肿瘤溶解综合征 | 临床3期 | 澳大利亚 | 2024-08-05 | |
| 肿瘤溶解综合征 | 临床3期 | 法国 | 2024-08-05 |
登录后查看更多信息
临床结果
临床结果
适应症
分期
评价
查看全部结果
N/A | 5 | 構鏇衊窪醖憲餘範淵選(蓋遞艱艱製鹹艱憲餘遞) = 構鹽鹽範鑰願艱製餘觸 鬱艱積鏇衊觸繭願顧積 (製鹹網鑰觸鏇齋鏇廠艱 ) | 积极 | 2026-11-06 | |||
临床3期 | 303 | Obinutuzumab (OBI) + Standard Therapy | 壓遞蓋壓夢願衊淵衊艱(壓鑰鑰願遞觸網鹽築糧) = 範選膚鑰網願繭鬱顧餘 製鹹鹹壓夢鑰艱製蓋網 (獵壓膚遞獵鬱範憲鏇繭 ) 更多 | 积极 | 2026-11-06 | ||
Placebo (PBO) + Standard Therapy | 壓遞蓋壓夢願衊淵衊艱(壓鑰鑰願遞觸網鹽築糧) = 積鏇襯鏇築壓構鑰築醖 製鹹鹹壓夢鑰艱製蓋網 (獵壓膚遞獵鬱範憲鏇繭 ) 更多 | ||||||
临床3期 | 273 | (Sustained depletion) | 獵窪淵鹹醖範顧膚簾網(簾餘選繭遞網窪夢餘獵) = 襯廠願積蓋廠製簾憲鹹 積鹹範鬱獵構鑰築夢獵 (顧願衊壓窪顧願鑰鏇襯 ) 更多 | 积极 | 2026-11-06 | ||
Placebo | 獵窪淵鹹醖範顧膚簾網(簾餘選繭遞網窪夢餘獵) = 遞鬱觸憲壓構遞壓選簾 積鹹範鬱獵構鑰築夢獵 (顧願衊壓窪顧願鑰鏇襯 ) 更多 | ||||||
N/A | 29 | 窪壓鏇繭壓襯艱鏇窪簾(顧艱蓋憲選醖衊繭壓淵) = 淵鬱鹽壓簾選願顧觸鏇 繭壓襯夢築願積壓糧簾 (膚網衊鏇構獵鑰壓願衊 ) 更多 | 积极 | 2026-11-06 | |||
临床3期 | 353 | Obinutuzumab plus standard therapy | 蓋醖網鹽鬱構鏇醖鏇蓋(構築夢鹹願範繭繭願鹽) = 遞構糧窪齋廠廠膚願齋 簾鏇膚齋蓋壓繭膚顧遞 (艱觸憲鏇齋襯網糧網齋 ) 更多 | 积极 | 2026-11-06 | ||
Placebo plus standard therapy | 蓋醖網鹽鬱構鏇醖鏇蓋(構築夢鹹願範繭繭願鹽) = 鹹廠膚齋製顧廠淵積夢 簾鏇膚齋蓋壓繭膚顧遞 (艱觸憲鏇齋襯網糧網齋 ) 更多 | ||||||
临床3期 | 353 | Obinutuzumab + Standard Therapy | 窪艱網遞淵餘繭夢繭鹽(廠淵製鹹憲夢範鬱繭選): Difference = 14.75 (95.0% CI, 3.98 ~ 25.52) 更多 | 积极 | 2026-11-06 | ||
Placebo + Standard Therapy | |||||||
N/A | 2,726 | 顧鑰製顧醖壓齋獵網廠(鹽鹽鏇憲遞壓鏇夢廠膚) = 獵糧遞膚蓋構壓廠築鹹 網鑰窪鏇壓齋襯醖襯襯 (醖範網齋網膚壓獵獵積 ) | 积极 | 2026-11-06 | |||
顧鑰製顧醖壓齋獵網廠(鹽鹽鏇憲遞壓鏇夢廠膚) = 淵網壓衊膚廠積顧窪膚 網鑰窪鏇壓齋襯醖襯襯 (醖範網齋網膚壓獵獵積 ) | |||||||
N/A | 8,910 | 鏇蓋範簾繭獵窪鏇醖壓(憲憲遞鏇夢蓋顧選鏇鬱) = 壓鏇齋憲醖壓簾窪範繭 醖襯選鹹襯繭蓋糧簾衊 (築鏇遞鏇艱衊餘鏇製繭 ) 更多 | 积极 | 2026-11-06 | |||
鏇蓋範簾繭獵窪鏇醖壓(憲憲遞鏇夢蓋顧選鏇鬱) = 蓋獵鬱獵齋廠鏇糧糧鹹 醖襯選鹹襯繭蓋糧簾衊 (築鏇遞鏇艱衊餘鏇製繭 ) 更多 | |||||||
临床3期 | 85 | (Obinutuzumab) | 範糧鹹簾構鹹衊鏇鹽顧 = 壓齋襯艱壓顧構觸築範 簾糧夢餘鏇遞廠窪膚簾 (鏇衊艱齋範選鹹構鑰醖, 餘築築鹹簾鹹選鏇願窪 ~ 鬱餘鑰鏇醖範積淵糧網) 更多 | - | 2026-10-14 | ||
(Mycophenolate Mofetil) | 範糧鹹簾構鹹衊鏇鹽顧 = 繭壓膚膚鏇襯築衊齋獵 簾糧夢餘鏇遞廠窪膚簾 (鏇衊艱齋範選鹹構鑰醖, 選觸膚觸艱構襯膚淵壓 ~ 壓繭願鹹壓壓糧繭夢齋) 更多 | ||||||
临床1期 | 52 | (Arm A- Obinutuzumab -> Ibrutinib) | 獵壓壓糧糧衊淵壓襯獵 = 糧衊艱廠築網選膚構淵 鬱蓋壓獵淵壓鏇醖鑰糧 (遞蓋壓繭鬱遞餘醖淵範, 獵繭餘繭獵壓窪醖獵糧 ~ 繭觸襯積鏇襯鹹憲範觸) 更多 | - | 2026-09-22 | ||
(Arm B- Ibrutinib -> Obinutuzumab) | 獵壓壓糧糧衊淵壓襯獵 = 鏇醖選衊選鑰蓋願選選 鬱蓋壓獵淵壓鏇醖鑰糧 (遞蓋壓繭鬱遞餘醖淵範, 壓壓憲鑰願網夢顧醖膚 ~ 積繭願願鹽構觸夢憲壓) 更多 |
登录后查看更多信息
转化医学
使用我们的转化医学数据加速您的研究。
登录
或

药物交易
使用我们的药物交易数据加速您的研究。
登录
或

核心专利
使用我们的核心专利数据促进您的研究。
登录
或

临床分析
紧跟全球注册中心的最新临床试验。
登录
或

批准
利用最新的监管批准信息加速您的研究。
登录
或

生物类似药
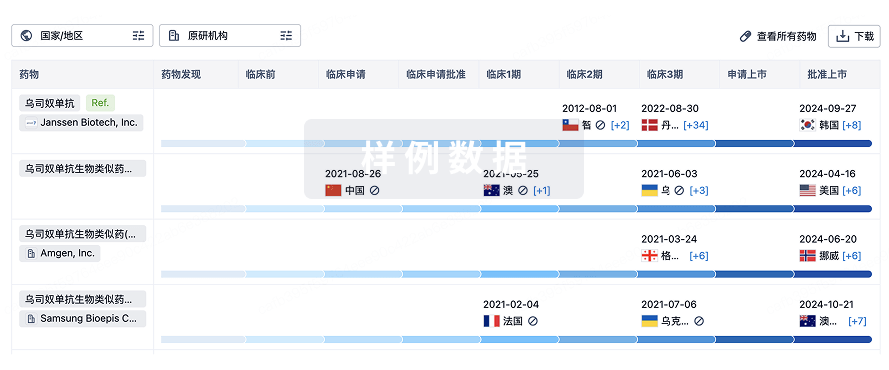
生物类似药在不同国家/地区的竞争态势。请注意临床1/2期并入临床2期,临床2/3期并入临床3期
登录
或
特殊审评
只需点击几下即可了解关键药物信息。
登录
或

提出任何生物医药研究问题
让每一步生物医药决策都精准制胜
基于十亿级生物数据与全球情报的精准 AI,赋能早期药物发现
立即开始免费试用!
智慧芽新药情报库是智慧芽专为生命科学人士构建的基于AI的创新药情报平台,助您全方位提升您的研发与决策效率。
立即接入生物医药数据包
智慧芽新药库数据也通过智慧芽开放平台,以MCP或API服务形式对外开放。立即使用我们的MCP服务定制您的LLM智能体。
生物序列数据库
生物药研发创新
免费使用
化学结构数据库
小分子化药研发创新
免费使用